


Contract manufactures are faced with multiple challenges when determining whether to implement process analytical technology into their clients' or their own infrastructure.
Contract manufactures are faced with multiple challenges when determining whether to implement process analytical technology into their clients' or their own infrastructure.
Pharmaceutical Technology Europe
To produce an application or solution for a specific domain, a vendor must demonstrate a thorough understanding of the domain and its specific challenges.
Pharmaceutical Technology Europe
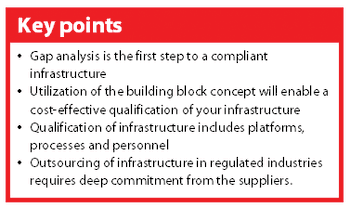
A new Good Automated Manufacturing Practice (GAMP) guide on IT Infrastructure Control & Compliance was launched in Chicago (IL, USA) 23 August 2005.1 The guide is intended to support pharmaceutical companies in their effort to establish a well-defined and compliant infrastructure. This article discusses different aspects of the guide that may support your organization in getting — and keeping — your infrastructure under control.
ePT--the Electronic Newsletter of Pharmaceutical Technology
The process analytical technology (PAT) initiative has been percolating at the US Food and Drug Administration for a long time, explained FDA's John E. Simmons at the AAPS Annual Meeting and Exposition on Wednesday. "If you think of PAT as an isolated set of applications, I think you are missing the point," Simmons said. "The FDA would like PAT to become commonplace?not to be an initiative, but common practice."
ePT--the Electronic Newsletter of Pharmaceutical Technology
As a pharmaceutical formulation tool, molecular simulation is currently in its early infancy. Nonetheless, presenters at Wednesday?s AAPS Annual Meeting and Exposition demonstrated that the technology is beginning to attract some interest. The topic was discussed in a presentation titled "Application of Molecular Simulations to Formulation Development and Stability Prediction."
ePT--the Electronic Newsletter of Pharmaceutical Technology
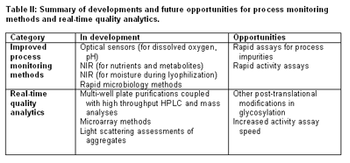
"The better we understand the relationship between process parameters and product attributes, the better control we'll have over product quality," said Beth Fowler, PhD, during Tuesday?s session on process monitoring at the AAPS Annual Meeting.
ePT--the Electronic Newsletter of Pharmaceutical Technology
Pressures to save API are driving formulation developers toward smaller-scale laboratory processes, while pressures to save time put a premium on more- accurate laboratory scale tools.
New FDA Guidance on Electronic Common Technical Document Submissions
Coordinating validation efforts throughout an organization requires an accurate and timely overview and a validation master plan (VMP).
To implement process analytical technology systems into the current information technology landscape, manufacturers will need to adopt continuously available systems and infrastructures.
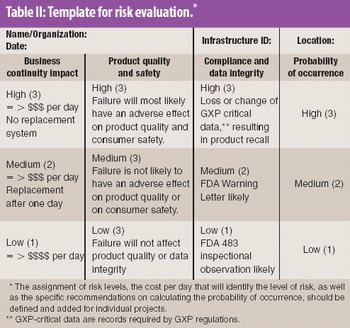
Risk analysis and evaluation of software and computer systems is a good tool to optimize validation costs by focusing on systems with high impact on both the business and compliance.
Pharmaceutical Technology Europe
Regulatory and compliance requirements are frequently changing, and if that wasn't enough there is also numerous varying regulations across products, services and countries.
In 1987, when the US Food and Drug Administration issued its Guideline on General Principles of Process Validation, a young FDA reviewer asked her supervisors.
Pharmaceutical Technology Europe
Reading the good automated manufacturing practice (GAMP 4) guide acquaints you with the now classic and almost famous V-model.1 The V-model, originally used for describing a validation workflow of IT and automated systems, is easy to understand and very good at ensuring that the requirements and design are built into the final solution. It is also extremely versatile and can be used for almost any type of validation task you could meet in a development phase.
A good electronic records managment program will not just impress regulators. It also can boost profitability.
This article describes the Part 11 requirements that companies must address when automating their quality management processes and offers practical tips for compliance.
Glossary of computer and software terms.
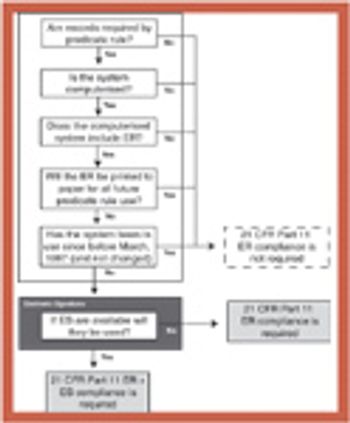
Nearly eight years have elapsed since the US Food and Drug Administration's 21 CFR Part 11 regulations on the use of electronic records and electronic signatures went into effect (1). In Sept. 2003, FDA issued a guidance document covering the scope and application of Part 11, which described how the agency intends to interpret and enforce the requirements during its ongoing re-examination of the regulations (2). Many in the pharmaceutical industry view the guidance as a positive development that will lead to a simplified FDA approach to Part 11 and a significant reduction in the industry's compliance burden. But this shift in FDA's interpretation and its intended use of enforcement discretion has not ended the controversy and confusion surrounding Part 11 and its requirements.
The confusion surrounding the application of 21 CFR Part 11 can be solved by focusing on the common rationale that both protects the public health and drives the fundamental basis of the industry's business: good science.
The use of quality management software (QMS) to automate manufacturing quality processes quickly is becoming an industry-wide initiative. Companies are turning to commercial off-the-shelf systems (COTS) to simplify implementation and validation efforts. Regardless of the system selected, the automation of these critical quality processes is subject to electronic records and electronic signatures (ER/ES) requirements, as set forth in the 21 CFR Part 11 regulation (1). Therefore, companies must adopt a comprehensive but manageable approach to Part 11 compliance as they begin to automate the processes that support product quality and efficacy to ensure they meet the requirements of good manufacturing practices and all predicate rules.
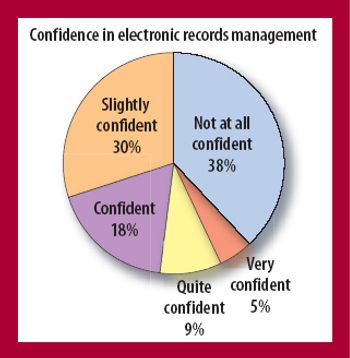
Pharmaceutical firms have every reason to feel confused and even a bit bewildered by what the US Food and Drug Administration expects from them when it comes to controlling electronic records. At times over the past nine years it has been a bit like riding a roller coaster that you weren't allowed to get off.
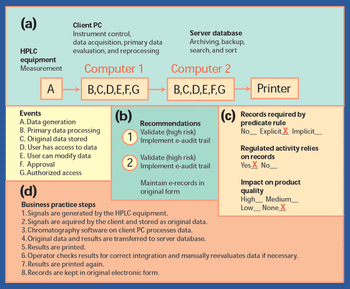
In 1997, the US Food and Drug Administration issued a regulation"Rule 21 CFR Part 11," that provides criteria for the acceptance of electronic records, electronic signatures, and handwritten signatures (1). FDA issued the guidance in response to requests from the industry. With this regulation, electronic records can be equivalent to paper records and handwritten signatures. The rule applies to all industry segments regulated by FDA that include good laboratory practice, good clinical practice, and current good manufacturing practice.
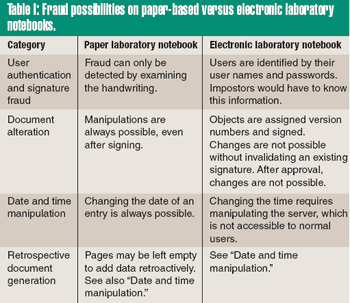
Paper notebooks are the accepted method for recording laboratory data and the ideas generated from that information in the pharmaceutical, biotech, and chemical industries. Nonetheless, the revolution in digital data processing has improved the way data is created, organized, and managed electronically, whether in the form of analytical data, images, documents, or multimedia files. The preservation of such information into a digital form offers the potential for online storage and retrieval, efficient search processes, and worldwide data transmission.
Purdue Pharmaceutical Manufacturing and Education Center Opens its Doors
FDA is finalizing guidances and enforcing compliance with manufacturing standards as part of its efforts to ensure drug safety.